What Happens When You Document But Don't Validate?
- A biotech company had everything in place for launch. Regulatory release documentation was complete. Inventory was packaged and ready. Retailers had confirmed shelf space. The timeline was locked. The product was approved. Everyone was ready to ship.
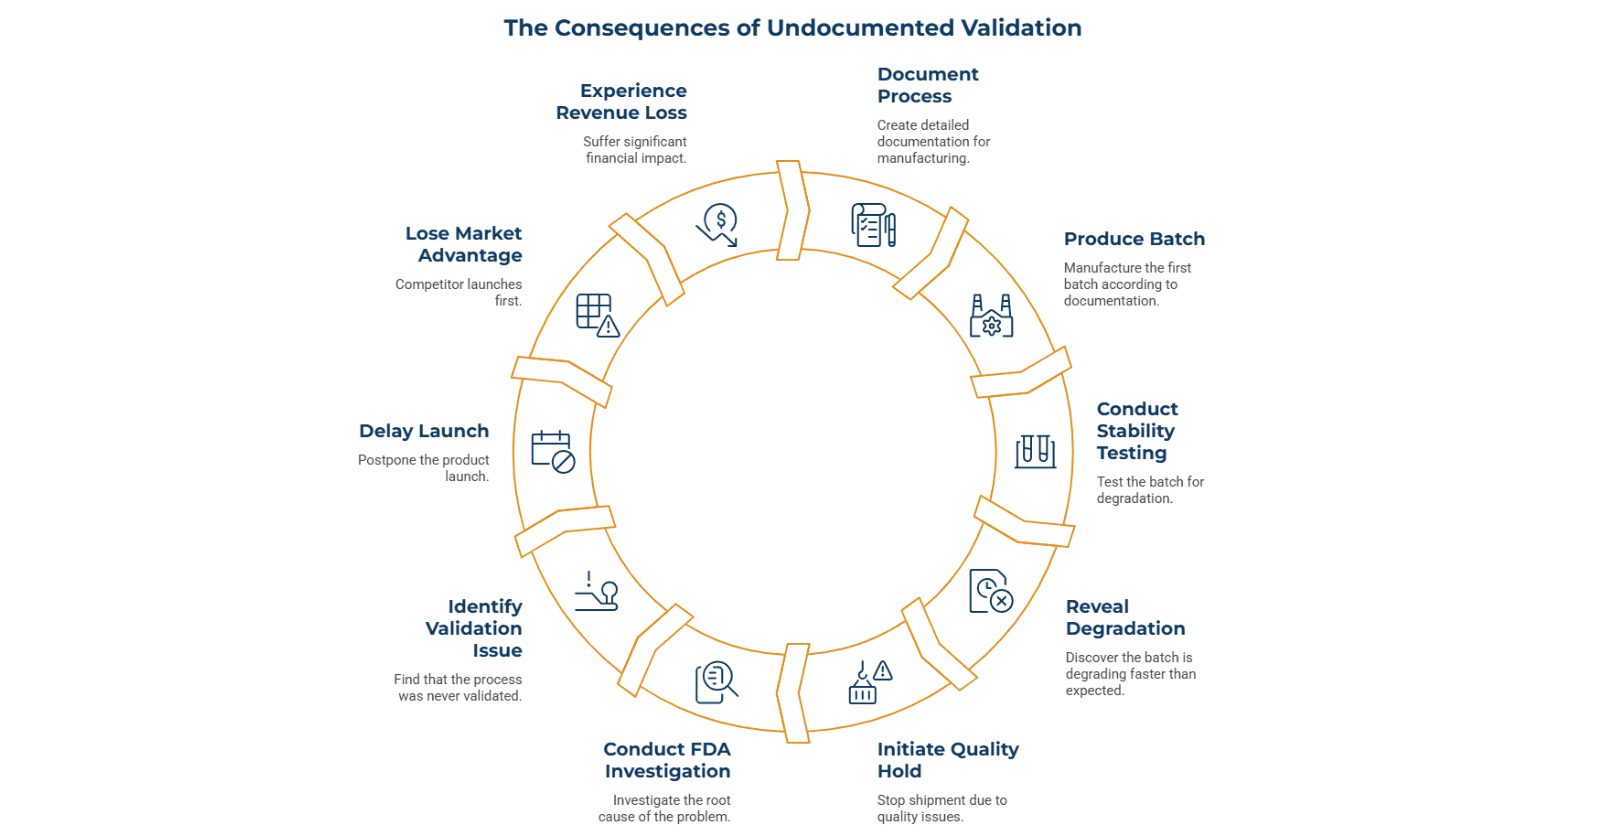
- Day 1 of commercial shipment: CDMO produced the first batch. It looked perfect. Documentation was complete. The process had been followed exactly as written. Then Week 2: stability testing revealed the batch was degrading faster than expected. Quality hold. The product couldn't ship.
- Six months later, an FDA investigation found the real problem. The manufacturing process was never properly validated during pre-production. The CDMO had followed the written procedure. But the procedure had never been scientifically proven to work consistently at commercial scale.
- Launch was delayed 8 months. First-mover advantage lost to a competitor who got to market first. The biotech company's revenue impact: $18 million in lost first-year sales.
- The process was documented. The batch looked good. But it wasn't validated. And there's a massive difference.
The False Confidence of Documentation
- Documentation feels like readiness. It shouldn't, but it does.
2. A biotech company and their CDMO create a Standard Operating Procedure (SOP). The procedure details every step: temperature set points, mixing times, hold periods, quality checks. Everything is written. Everything is documented. Everything looks professional.
3. The CDMO executes the procedure on small trial batches of maybe 100 units for testing purposes. Everything works. The batches meet specifications. The process follows the SOP perfectly. Quality signs off. R&D confirms results. Everyone agrees: the process works.
4. "Ready to launch," the team declares. Documentation is complete. The process has been executed. What could go wrong? Everything. Because documentation is not validation. Executing a procedure on 100-unit trial batches is not proof the procedure works on 500K-unit commercial batches.
5. But most teams don't realize this until it's too late.
Why Validation and Documentation are Not the Same
This is where the gap appears.
Documentation is: here's how we're going to make this product. Validation is: we've scientifically proven this process works consistently.
Those are completely different things. A documented process can fail. A validated process should work. The FDA knows the difference. They require validation. They audit it. They cite companies that skip it.
The biotech company had documentation. They didn't have validation. Here's what they were missing:
Parameter Sensitivity Testing:
Does the process care if temperature varies by 1 degree? What if mixing time is 5 minutes instead of 7? What if the hold period shifts? These aren't theoretical questions. They determine whether your process is robust or fragile. Parameter sensitivity testing answers them. The company never did it.
Batch-to-Batch Variability Characterization:
Manufacturing isn't perfectly consistent. One batch might have slightly different raw material properties. Equipment might behave differently. Operators might interpret timing slightly differently. A validated process knows how much variation it can tolerate. This company's process was untested against real-world variation.
Failure Mode Analysis:
What breaks the process? What causes batches to fail? What conditions destroy product quality? A validated process has tested these questions intentionally. This company had never asked them. They discovered the answers during commercial production. Too late.
Scale-Up Science:
A 100-unit batch and a 500K-unit batch behave differently. Temperature distribution isn't uniform in large vessels. Mixing patterns change. Hold time impacts are different. Validated processes account for scale. This company assumed small-scale results would translate directly. They didn't.
The Scale Gap Nobody Tests
- The most dangerous assumption in biotech manufacturing: small-scale success means large-scale success.
- It doesn't.
- Trial batches are 100 units. Commercial batches are 500K units. That's a 5,000x difference. The physics changes. The chemistry changes. The risks change.
- In the biotech company's case, the CDMO's mixing tanks behaved differently at large scale. The temperature distribution wasn't uniform. The product wasn't mixing evenly. Some portions of the batch were getting longer hold times than others. This created degradation that the small batches never revealed.
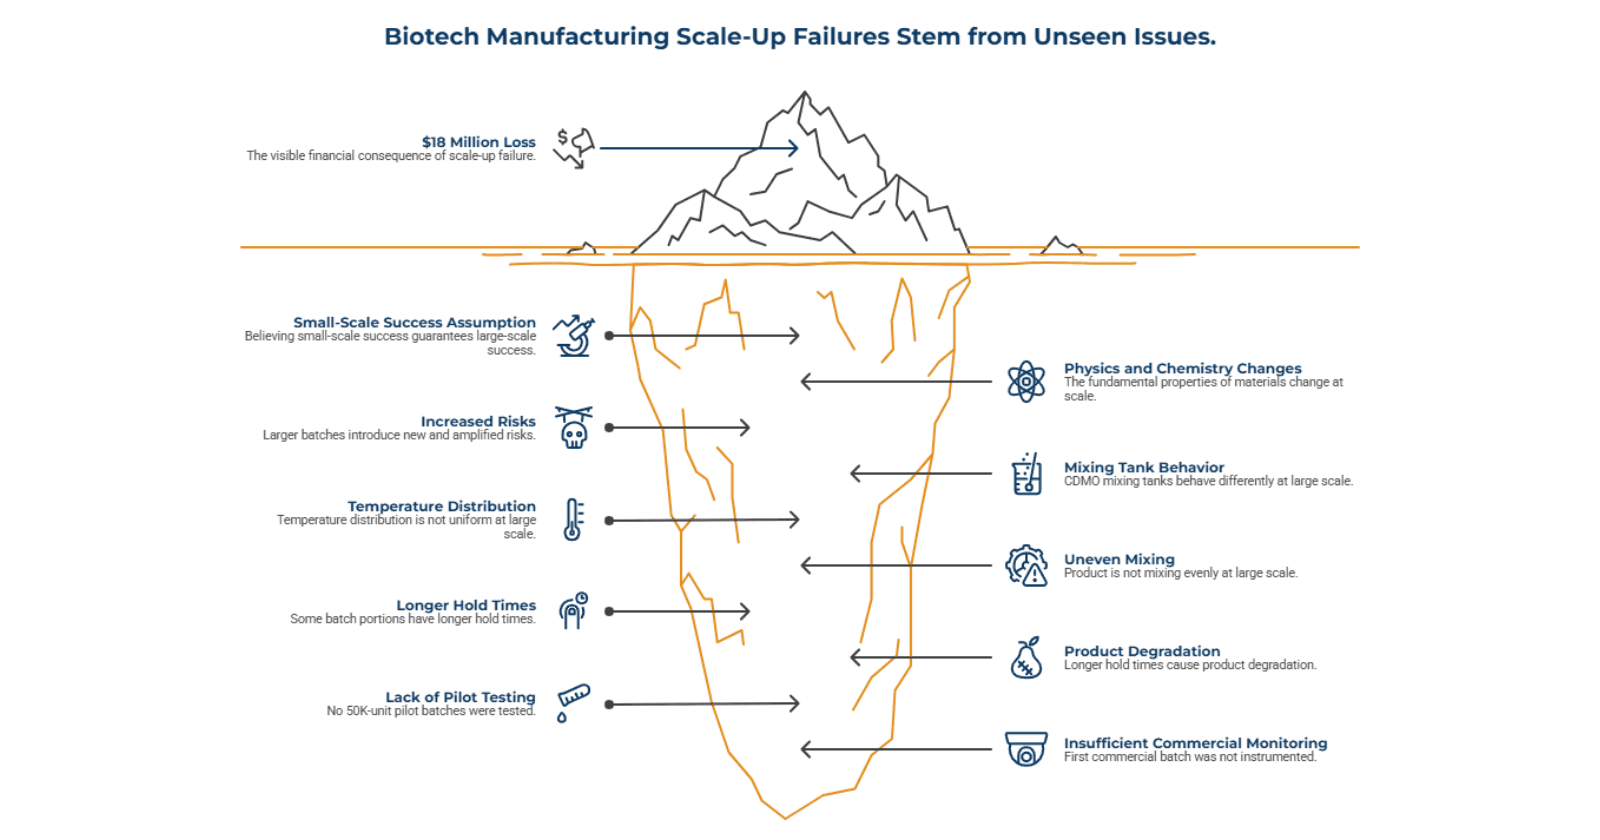
- Nobody had tested this. Nobody had intentionally made 50K-unit pilot batches to see if the process scaled. Nobody had instrumented a commercial batch with extra monitoring to catch problems. They assumed: we made 100 units successfully, so 500K units will work the same way.
- This assumption cost them $18 million.
- A validated process works backward from commercial scale. It tests at pilot scale first (50% of commercial size). It instruments the first commercial batch with extra monitoring. It doesn't assume. It proves.
How Governance Fails Before Launch
- The real problem wasn't the manufacturing. It was the governance.
2. Nobody owned the validation decision. R&D team knew validation was required. They mentioned it. But they weren't responsible for launch timeline. Manufacturing team wanted to launch. Validation felt like a delay. Quality team had audit responsibility but not decision authority. Nobody could say: "We're not launching until validation is complete."
3. This is the governance gap. When multiple departments own pieces of the decision but nobody owns the decision itself, validation gets skipped.
4. In this company:
The launch decision got made based on timeline pressure, not readiness. And when the first batch failed, everyone was surprised.
When FDA Finds What You Missed
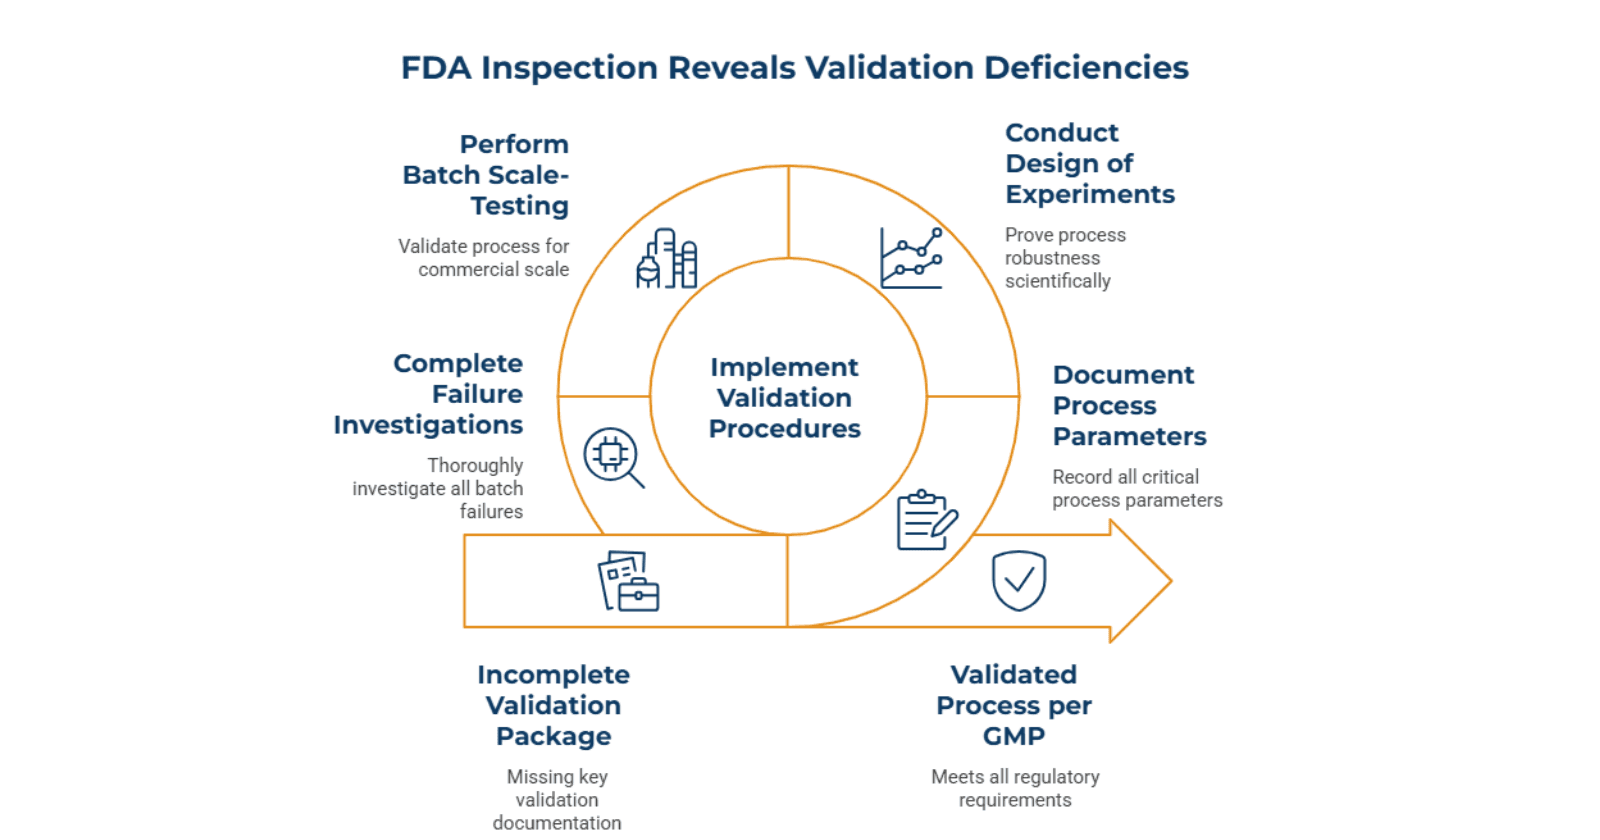
- Six months after launch, FDA showed up for inspection. They wanted to see the validation package. Process parameter documentation. Design of Experiments proving robustness. Batch scale-testing data. Failure mode analysis.
- The company didn't have any of it.
- What they had: documentation of the procedure. Results from 100-unit trial batches. A failed commercial batch with incomplete failure investigation. No evidence that the process had been scientifically validated for commercial scale.
- FDA's finding: Process not validated per GMP requirements.
- The impact: import alert. Market withdrawal. Reputation damage. Revenue loss that extended far beyond the first product.
- This isn't theoretical. This is what happens when companies confuse documentation with validation.
What Visibility Actually Requires
Validation readiness requires three things companies typically don't implement.
First: Scientific Parameter Definition
Before manufacturing starts, identify the critical parameters. Temperature. Mixing time. Hold period. pH. All the factors that matter. For each one, run a sensitivity analysis: if this parameter drifts, what happens? Does quality suffer? By how much? Define operating ranges based on science, not guesswork.
This takes time. It requires design of experiments. It costs money. But it proves the process is robust before commercial production.
2. Second: Scale-Up Testing Before Launch
Small-scale batches don't reveal large-scale problems. Test at pilot scale (50% of commercial size). Instrument the first commercial batch with extra monitoring. Document everything. Don't assume scale works. Prove it.
3. Third: Governance Authority
Assign someone to own the "no launch until validated" decision. Give them veto power. Make it clear: validation must be complete before commercial production starts. Timeline pressure cannot override this requirement.
What Actually Works
- Companies that avoid validation failures implement a structured pre-launch gate.
- One biotech client did this right. Here's what changed their outcome.
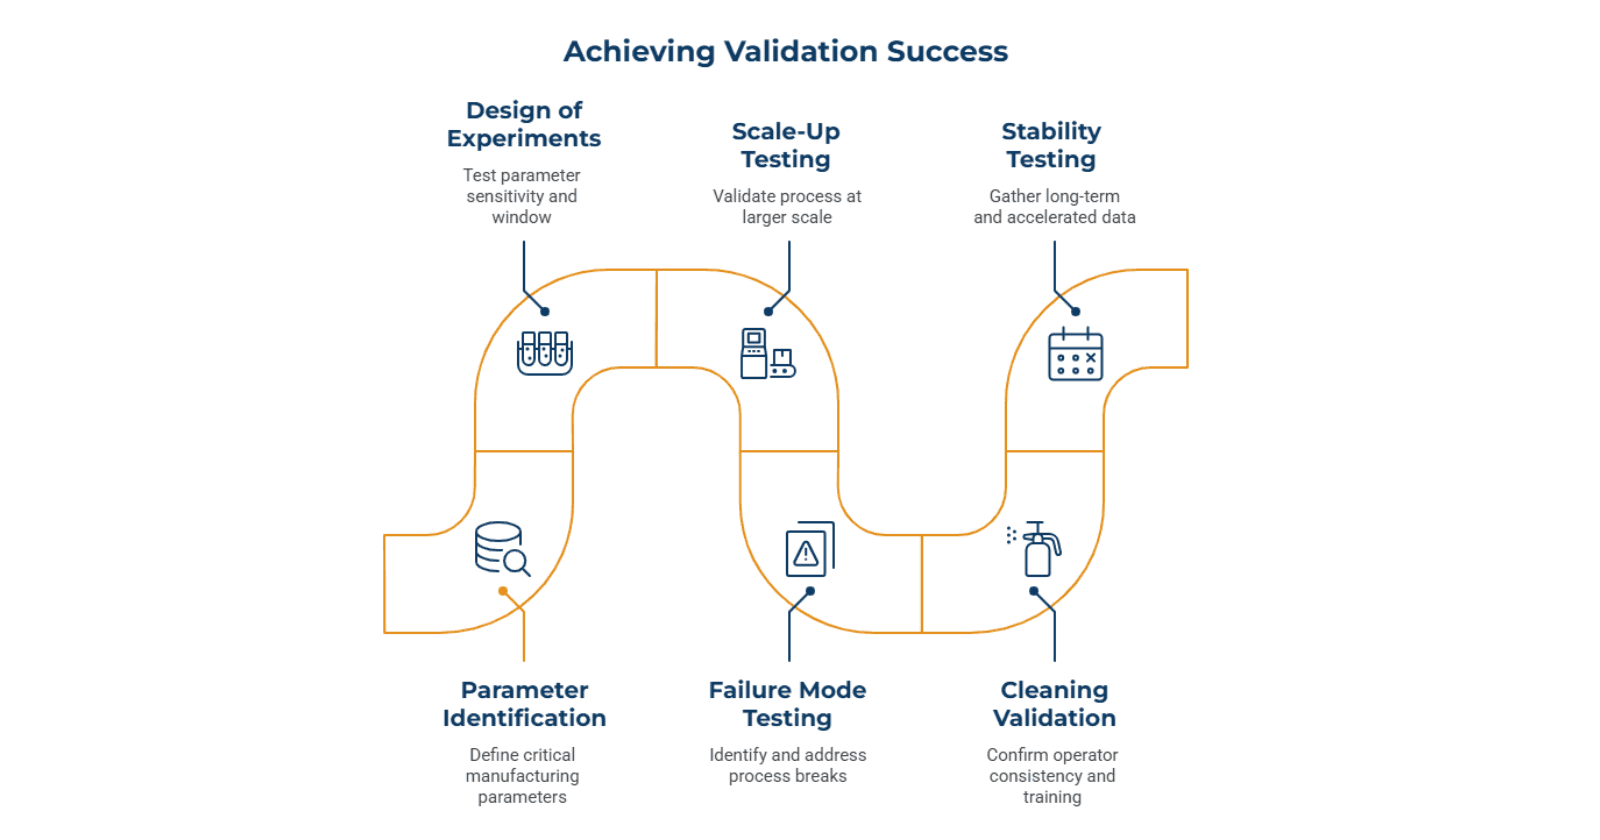
- They identified critical parameters before manufacturing: temperature (±2°C), mixing time (±10%), hold period (±5%), final pH (±0.5 units). Then they ran the design of experiments to test sensitivity. They made 10-unit pilot batches varying each parameter. They mapped the operating window.
- They made 50K-unit pilot batches to test scale-up. They instrumented the first commercial batch with extra sensors to monitor temperature distribution, mixing patterns, and hold time impacts. Results were documented and analyzed.
- They tested failure modes intentionally. What breaks this process? They found that product degradation accelerated above 25°C. They found that uneven mixing created hot spots. They found that batch-to-batch consistency required operator training on mixing technique.
- They validated cleaning between batches. They confirmed operator variation didn't cause quality issues with proper training.
- They ran stability testing: long-term data at shelf-life conditions. Accelerated data under stress. They had shelf-life data before launch, not discovered during commercial production.
- Governance: Quality VP had veto authority. Launch was approved only after validation checklist was complete: Timeline pressure mattered. But not more than validation completion. Result: Process validated before launch. Scale risks identified and managed before commercial production. FDA inspection found complete documentation. The launch happened on schedule. No surprises.
The Insight
Documentation is not validation. This distinction separates companies that launch successfully from companies that crash during commercial production. Validation is science. It's testing. It's proof. It's intentionally breaking the process to understand its limits. It takes time. It costs money. But it prevents $18 million revenue losses.
The companies that succeed don't skip validation. They plan for it. They start validation work months before commercial launch. They make governance decisions clear: validation must be complete before manufacturing starts.
Companies that confuse documentation with validation discover the gap at the worst possible moment when FDA audits, when batches fail, when revenue is already lost.The choice is simple. Do validation before launch or discover validation gaps after. One costs three months and $500K. The other costs $18 million and your market position.
JASPAUL
I'm Jaspaul, Operational Review Specialist at Gyan Solutions. With 6+ years in pharmaceutical supply chain consulting, I help biotech, CDMO, and medical device leaders build visible, resilient supply chains through business automation and operational alignment.